Electronic-Structure Methods for Materials Science - Advanced Quantum ESPRESSO Tutorial
Explore advanced electronic-structure methods for materials science, focusing on Hubbard and Koopmans functionals from linear response in Quantum ESPRESSO.
Electronic and Optical Spectra from Optimally-Tuned Functionals - QE Tutorial 2022
Explore optimally-tuned functionals for accurate electronic and optical spectra calculations in advanced quantum systems.
Exchange-Correlation Functionals of Density Functional Theory - QE School 2023 Lecture 1.3
Explore advanced exchange-correlation functionals in DFT, focusing on Hubbard and Koopmans functionals from linear response for enhanced quantum calculations.
Maximally Localized Wannier Functions: Theory and Recent Developments - Lecture 1.4
Explore maximally localized Wannier functions, their theoretical foundations, and latest advancements in quantum materials research.
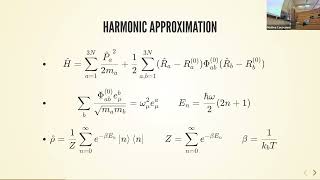
Beyond Harmonic Phonons: Phase Diagrams and Phase Transitions - QE School 2023 Lecture 2.3
Explore advanced quantum mechanics concepts, focusing on phase diagrams and transitions beyond harmonic phonons in materials science.
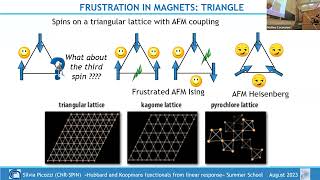
Successes and Failures of DFT+U in Ferroic Materials - Lecture 3.4
Explore successes and failures of DFT+U in ferroic materials, focusing on Hubbard and Koopmans functionals from linear response in advanced quantum calculations.
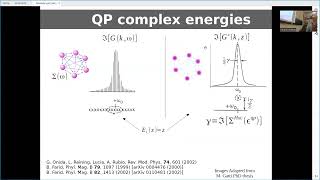
Ab Initio Many-Body Perturbation Theory - QE School 2023 Lecture 4.5
Explore advanced quantum mechanics with a focus on Hubbard and Koopmans functionals, delving into linear response theory and many-body perturbation techniques.
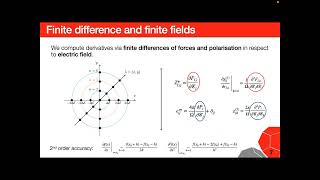
AiiDA-Vibroscopy: All-Functional Infrared and Raman Spectra - Lecture 5.2
Explore AiiDA-Vibroscopy for calculating infrared and Raman spectra using various functionals in Quantum ESPRESSO, enhancing materials research capabilities.
Computational Workflows for an Accelerated Design of Novel Materials - QE School 2023 Lecture 5.3
Explore computational workflows for accelerated design of novel materials, focusing on Hubbard and Koopmans functionals from linear response in advanced quantum calculations.
Spin-Orbit Related Phenomena in Magnets from Density Functional Theory
Explore spin-orbit physics in magnets using density functional theory. Understand electronic structure changes, magnetic interactions, and their impact on condensed matter phenomena.
Recent Advances in Density Functional Theory for Complex Correlated Materials
Explore recent advances in density functional theory for complex correlated materials, presented by Dr. Christopher Lane of Los Alamos National Lab.
Crystallization Machine Learning - Theory and Simulation Techniques
Explore machine learning applications in crystallization, from molecular dynamics to enhanced sampling techniques for rare events and homogeneous nucleation.
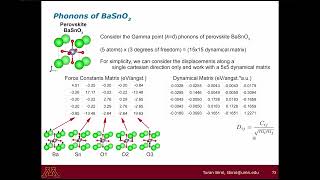
Phonons and Structural Transitions from DFT and Group Theory - Part 2
Explore phonons and structural transitions using DFT and group theory, advancing understanding of material properties and phase changes.
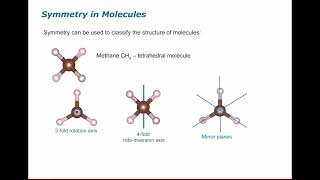
Phonons and Structural Transitions from DFT and Group Theory - Part 1
Explore phonons and structural transitions using DFT and group theory, gaining insights into fundamental concepts and advanced computational techniques.
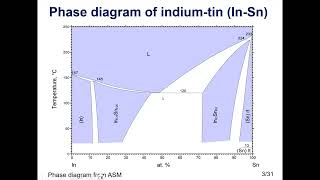
First Principles Thermodynamics
Explore computational condensed matter using first principles thermodynamics, gaining insights into advanced techniques and applications.
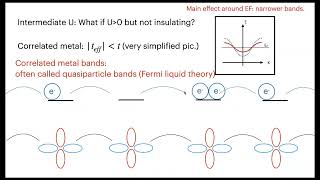
Concepts in the Computational Study of Correlated Electron Materials
Explore computational methods for studying correlated electron materials, focusing on advanced concepts and techniques in materials science and chemistry.
Facilitating Electronic Structure Computations on GPU-based Exascale Platforms
Explore GPU-based electronic structure computations for exascale platforms using PROGRESS and BML libraries. Learn implementation strategies, performance optimization, and scaling techniques.
CHGNet: A Pre-trained Universal Interatomic Potential for Electron-Coupled Ionic Systems
Explore CHGNet, a graph-neural-network-based machine-learning potential for modeling universal potential energy surfaces in large-scale simulations with complex electron interactions.
Efficient Calculations of Electronic Structures with Machine-Learning Models
Explore machine learning models for efficient electronic structure calculations in materials discovery and drug design, focusing on density functional theory and the Materials Learning Algorithms library.
Disordered Hyperuniformity in Two-Dimensional Materials
Explore disordered hyperuniformity in 2D materials, its impact on electronic structures, and role of topological defects in preserving hyperuniformity in amorphous materials.
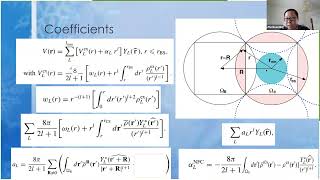
Shaped Solution to the Poisson Equation in the Full-Potential KKR Method
Explore a practical and accurate solution to the Poisson equation in full-potential KKR, focusing on the removed-sphere method for disordered materials.