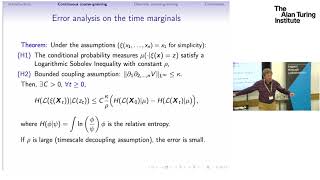
Coarse-Graining Stochastic Dynamics
Explore techniques for simplifying complex stochastic systems through coarse-graining, enabling practical computation and accurate prediction in various scientific fields.
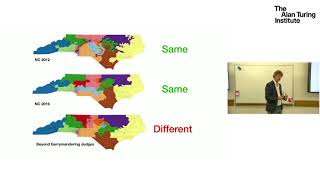
Understanding the Geopolitical Structure of the USA
Explore statistical approaches for analyzing geopolitical structures in the USA, combining data analysis with complex systems modeling to uncover crucial features and patterns.
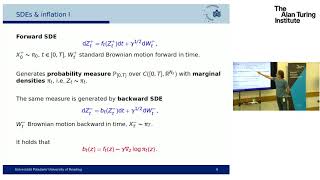
Learning Models by Making Them Interact
Explore new approaches to data-driven modeling of complex systems, combining statistical inference, high-throughput computation, and physical laws for more accurate predictions in various scientific fields.
The Complexity of Fermions in Quantum Information and Beyond
Exploring fermion simulation challenges in quantum computing, highlighting new data structures and complexity results for ground state problems in chemistry, physics, and computer science.
Electronic Structure Theory via Classical and Quantum Computation - IPAM at UCLA
Explore electronic structure theory through classical and quantum computation, focusing on innovative techniques for modeling molecular systems and their potential applications.

Binding Affinity Prediction with Machine Learning-Based Docking - Lab 2
Explore ML-based docking techniques for predicting binding affinity in drug discovery, featuring insights from experts Stephan Thaler and Cristian Gabellini.
Exploring Molecular Space and Active Learning in Drug Discovery
Explore molecular space and active learning techniques for drug discovery with AI pioneer Yoshua Bengio's insights from the 2024 ML Summer School at Mila.
Lab 3 - De Novo Generation Explanation
Explore de novo molecule generation techniques with experts Emmanuel Bengio and Julien Roy in this recorded session from the 2024 Machine Learning for Drug Discovery Summer School.
Learning ML Interatomic Potentials - Day 2
Explore ML interatomic potentials for drug discovery with expert Gianni De Fabritiis. Gain insights into advanced techniques and applications in computational chemistry.
LLMs in Drug Discovery - Day 5
Explore the application of Large Language Models in drug discovery, focusing on innovative approaches and potential breakthroughs in pharmaceutical research.
Harnessing Geometric ML for Molecular Design - Day 3
Explore geometric machine learning techniques for innovative molecular design in drug discovery with expert Michael Bronstein.
Generative Models of Molecular Structures - Day 3
Explore generative models for molecular structures, focusing on applications in drug discovery and computational chemistry.
Machine Learning in Structure-Based Drug Discovery - Day 2
Explore machine learning applications in structure-based drug discovery, focusing on innovative techniques and their impact on pharmaceutical research.
Polaris - Industry-Led Initiative to Critically Assess ML for Real-World Drug Discovery
Explore an industry-led initiative critically assessing machine learning's impact on real-world drug discovery through the Polaris project.
Leveraging Molecular ML and Property Prediction in Drug Design
Explore cutting-edge molecular machine learning techniques for drug design, focusing on property prediction to enhance pharmaceutical development.
Flow Annealed Importance Sampling Bootstrap - Machine Learning for Complex Distributions
Explore Flow Annealed Importance Sampling Bootstrap for approximating complex target distributions. Learn about normalizing flows, annealed importance sampling, and applications to molecular systems.
Artificial Chemical Intelligence: AI for Chemistry and Chemistry for AI
Explore the integration of AI with chemistry, focusing on trustworthy frameworks for molecular discovery. Learn about innovative approaches to predict mechanisms and overcome challenges in computational chemistry.
Crippen's logP as a Quantitative Molecular Benchmark for Explainable AI Heatmaps
Explore Crippen's logP as a quantitative benchmark for explainable AI heatmaps in molecular modeling. Learn about atomic attribution methods and their applications in chemical property prediction.
MACE: Higher Order Equivariant Message Passing Neural Networks for Fast and Accurate Force Fields
Explore higher-order equivariant message passing neural networks for fast, accurate force fields in computational chemistry. Learn about MACE's innovative approach using four-body messages to enhance efficiency and accuracy.
Introducing Bioptic - Machine Learning for Drug Discovery
Explore Bioptic's architecture, training, and scaling for machine learning in drug discovery with Vlad Vinograv.
Molecular Machine Learning Conference 2023 - Opening Remarks
Explore opening remarks from Yoshua Bengio and Dominique Beaini at MoML 2023, covering conference highlights, tributes, and the future of molecular machine learning.