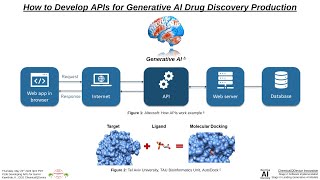
How to Develop APIs for Generative AI Drug Discovery Production
Explore platforms for creating APIs from Large Language Models to enhance drug discovery workflows. Learn to increase efficiency, reduce errors, and lower costs through synchronization and automation.
Data-Efficient Graph Grammar Learning for Molecular Generation
Explore data-efficient neurosymbolic generative models for molecular generation using learnable graph grammars, optimized for small datasets and physically synthesizable molecules.

Materials Discovery in Challenging Spaces with Machine Learning - Lecture 32
Explore materials discovery using machine learning, focusing on transition metal complexes and metal-organic frameworks with Prof. Heather Kulik from MIT.
Ab-initio Green's Functions Methods for Molecules and Solids - Lecture 36
Explore ab-initio Green's functions methods for molecules and solids, examining their accuracy and applications in computational materials science.
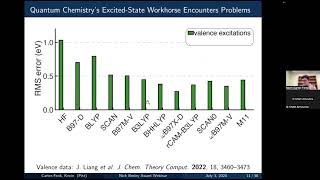
EA-TDDFT for Calculating X-ray Absorption Spectra
Explore EA-TDDFT for accurate X-ray absorption spectra calculations. Learn its applications in liquid-phase systems, glycerol aerosols, and Fe(CO)5 photodissociation, revealing insights into electronic and nuclear structure interplay.
Recent Advances in Density Functional Theory for Complex Correlated Materials
Explore recent advances in density functional theory for complex correlated materials, presented by Dr. Christopher Lane of Los Alamos National Lab.
Crystallization Machine Learning - Theory and Simulation Techniques
Explore machine learning applications in crystallization, from molecular dynamics to enhanced sampling techniques for rare events and homogeneous nucleation.
Concepts in the Computational Study of Correlated Electron Materials
Explore computational methods for studying correlated electron materials, focusing on advanced concepts and techniques in materials science and chemistry.
Facilitating Electronic Structure Computations on GPU-based Exascale Platforms
Explore GPU-based electronic structure computations for exascale platforms using PROGRESS and BML libraries. Learn implementation strategies, performance optimization, and scaling techniques.
CHGNet: A Pre-trained Universal Interatomic Potential for Electron-Coupled Ionic Systems
Explore CHGNet, a graph-neural-network-based machine-learning potential for modeling universal potential energy surfaces in large-scale simulations with complex electron interactions.
Efficient Calculations of Electronic Structures with Machine-Learning Models
Explore machine learning models for efficient electronic structure calculations in materials discovery and drug design, focusing on density functional theory and the Materials Learning Algorithms library.
Diffusion Models
Explore advanced concepts in diffusion models, including theory, applications, and recent developments in probabilistic AI.
Accelerating Drug Discovery with Generative AI
Discover how generative AI accelerates drug discovery through advanced machine learning techniques and innovative computational approaches in pharmaceutical research.
The Frontiers of Modeling Atoms with AI
Explore cutting-edge AI applications in atomic modeling, focusing on drug discovery and climate change solutions, with insights into machine learning potentials and renewable fuel synthesis catalysts.
Artificial Intelligence for Protein Modeling - DNA, RNA and Ion Docking Explained
Discover how AlphaFold3 revolutionizes structural biology through AI-powered protein modeling, enabling accurate predictions of 3D molecular structures, DNA/RNA interactions, and protein-protein binding mechanisms.
Generative AI for Drug Discovery Hit Identification and Optimization
Discover how generative AI revolutionizes drug discovery through innovative hit identification and optimization techniques, presented by a leading expert in machine learning research.
Machine Learning for Early Stage Drug Design - Property Prediction and Molecule Generation
Explore cutting-edge applications of machine learning in early-stage drug design, focusing on property prediction and molecule generation techniques for pharmaceutical development.
Structure-Based Virtual Ligand Screening with ICM-Pro
Master structure-based virtual ligand screening techniques using ICM-Pro software, from project setup and file preparation to running screens, analyzing results, and creating hitlists.
Learning Equivariant Non-Local Electron Density Functionals
Delve into advanced machine learning approaches for improving density functional theory through equivariant graph neural networks, focusing on non-local exchange-correlation functionals and molecular energy predictions.