Approaching Exact Quantum Chemistry by Semi-Stochastic and Selected-CI-Driven CC Computation
Explore advanced quantum chemistry techniques, including semi-stochastic methods and selected-CI-driven CC computation, for achieving near-exact results.
MRSF-TDDFT - A Simple Yet Powerful Quantum Theory
Explore a powerful quantum theory, MRSF-TDDFT, and its applications in computational chemistry and materials science.
Quantum Chemistry with Qiskit
Explore quantum chemistry applications using Qiskit, focusing on simulating molecular systems and solving electronic structure problems.
A Mathematical Approach to Coupled Cluster Methods
Explore mathematical foundations of Coupled Cluster methods for many-body problems, covering essential concepts and recent advances in quantum chemistry and nuclear physics.
Coupled-Cluster Theory for Ground and Excited Eigenstates
Explore advanced quantum computing concepts through coupled-cluster theory, examining ground and excited eigenstates and their applications in quantum systems.
Introduction to Applications of Quantum Computing to Quantum Chemistry - Class 1
Explore quantum computing applications in quantum chemistry, focusing on fundamental concepts and their practical implementation.
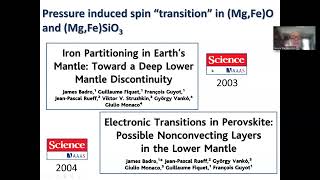
Iron Oxides and Silicates at Extreme Conditions - Advanced Quantum ESPRESSO Tutorial 2022
Explore iron oxides and silicates under extreme conditions, focusing on spin transitions and their impact on Earth's lower mantle seismology and dynamics.
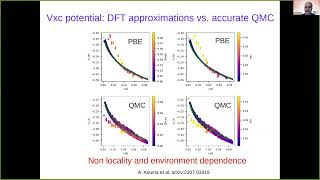
QE Tutorial 2022 - Design of Auxiliary Systems and Connectors for Spectroscopy
Explore advanced quantum mechanics concepts, focusing on Hubbard and Koopmans functionals in linear response, with emphasis on auxiliary systems and spectroscopy design.
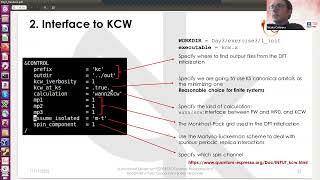
Spectral Properties of Finite and Extended Systems Using the KCW Code - QE Tutorial 2022
Explore spectral properties of finite and extended systems using KCW code in Quantum ESPRESSO, focusing on Hubbard and Koopmans functionals.
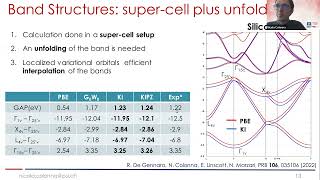
KCW: Implementation of Koopmans Functionals Based on DFPT - Advanced Quantum ESPRESSO Tutorial 2022
Explore Koopmans functionals implementation in Quantum ESPRESSO using density functional perturbation theory, enhancing electronic structure calculations.
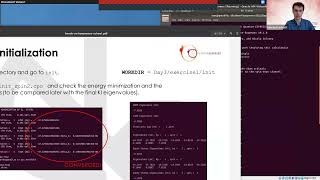
QE Tutorial 2022 - Spectral Properties of Finite and Extended Systems
Explore spectral properties of finite and extended systems through hands-on exercises using Quantum ESPRESSO, focusing on Hubbard and Koopmans functionals.
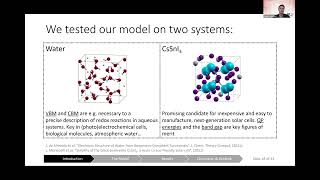
Accelerating the Calculation of Koopmans Screening Parameters Using Machine Learning
Accelerate Koopmans screening parameter calculations with machine learning techniques in Quantum ESPRESSO, enhancing efficiency in computational materials science.

Koopmans Functionals in Practice - QE Tutorial 2022
Explore Koopmans functionals in Quantum ESPRESSO, focusing on practical applications and advanced techniques for electronic structure calculations.
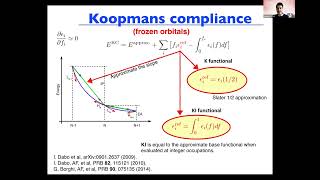
Koopmans Functionals and Spectral Potential in Quantum ESPRESSO - Tutorial 2
Explore Koopmans functionals in quantum mechanics, focusing on spectral potential connections and applications in materials science.
QE Tutorial 2022 - First-Principles Calculation of Hubbard Parameters
Explore first-principles Hubbard parameter calculations in quantum materials, focusing on chromium oxide and practical applications in advanced quantum simulations.
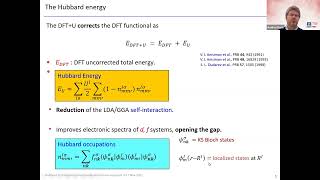
Phonons and Electron-Phonon Coupling Using DFPT+U - Advanced Quantum ESPRESSO Tutorial
Explore phonons and electron-phonon coupling using DFPT+U in Quantum ESPRESSO, focusing on advanced techniques for Hubbard and Koopmans functionals.
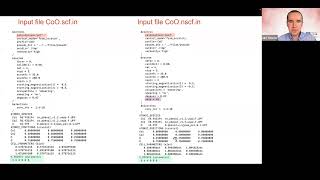
DFT+U and DFT+U+V: How Does It Work? - Hands-on Tutorial
Explore DFT+U and DFT+U+V methodologies in Quantum ESPRESSO through hands-on exercises, enhancing understanding of Hubbard-corrected density functional theory.
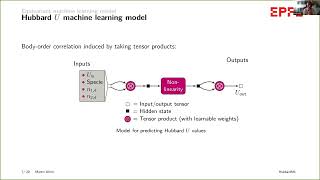
Accelerating the Calculation of Hubbard Parameters Using Machine Learning - QE Tutorial 2022
Learn advanced techniques for accelerating Hubbard parameter calculations in Quantum ESPRESSO using machine learning, with a focus on linear response methods and transferability.
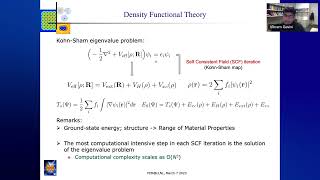
Fast, Accurate and Large-scale Ab-initio Calculations for Materials Modeling
Explore advanced computational methods for large-scale density functional theory calculations using adaptive finite-element discretization, enhancing materials modeling capabilities.
Competing Magnetisms and Onset of Static Magnetic Order in Sr2RuO4
Explore competing magnetic instabilities in Sr2RuO4, their link to superconductivity, and the onset of static magnetic order under strain using first-principles and self-consistent renormalization theories.
QC2: A Modular Open-Source Software for Quantum Chemistry
Explore QC2, a modular open-source software for quantum chemistry, its design principles, workflow, and applications in quantum computing and classical approaches.