DP-GEN: An Optimal Active Learning Protocol for Constructing Interatomic Potentials - Materials Simulation
Explore DP-GEN, an innovative active learning protocol revolutionizing molecular modeling by constructing accurate interatomic potentials. Learn its three-step iterative process and applications in materials simulation.
Physical Constraints on the Evolution of Viral Entry
Explore physical constraints on viral entry evolution with Dr. Peter Kasson. Gain insights into membrane dynamics, fusion mechanisms, and pandemic prediction through experimental and computational approaches.
Molly.jl - Molecular Dynamics in Julia
Explore Molly.jl for molecular dynamics simulations in Julia. Learn about its features, advantages, and applications in research, including differentiable molecular simulation and force field improvement.
Spin Orbit Coupling in QMCPACK
Explore spin orbit coupling implementation in QMCPACK with Cody Melton's workshop presentation, covering advanced quantum mechanics concepts and computational techniques.
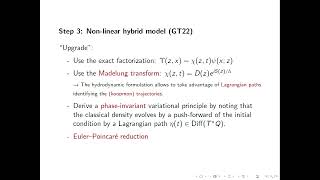
Madelung Transform and Variational Asymptotics in Born-Oppenheimer Molecular Dynamics
Explore variational asymptotics and Madelung transform in Born-Oppenheimer molecular dynamics, enhancing understanding of quantum chemistry principles.
Input Script for LAMMPS - Part 1
Learn to create and structure input scripts for LAMMPS molecular dynamics software, focusing on essential commands and parameters for simulation setup.
Basic Introduction to Molecular Dynamics
Explore molecular dynamics fundamentals, LAMMPS usage, visualization techniques, and key concepts like interatomic potentials and periodic boundary conditions.
Input Script for LAMMPS - Part 3
Learn to create and customize input scripts for LAMMPS molecular dynamics simulations, focusing on advanced techniques and best practices.
Input Script for LAMMPS - Part 2
Learn to create and customize input scripts for LAMMPS molecular dynamics simulations, enhancing your ability to model complex systems efficiently.
LAMMPS Exercises - Part 1
Explore LAMMPS molecular dynamics software through hands-on exercises, enhancing your understanding of simulation techniques and computational modeling.
LAMMPS Exercises - Part 5
Explore advanced LAMMPS exercises to enhance your molecular dynamics simulation skills and deepen your understanding of computational materials science.
Simulations of Multiphase Flow at Multiple Scales
Explore multiphase flow dynamics through simulations at various scales, from molecular to continuum, covering droplet behavior, emulsions, and fluid-solid interactions.
Computer Simulation of Water/Solid Interactions
Explore computer simulations of water-solid interactions, focusing on dynamic processes, structural features, and reactive applications in materials science.
Learning How to Build Molecules That Fold Like Proteins
Explore protein-like molecule folding with Prof. Michael Shirts, uncovering insights into building and understanding complex molecular structures.
Molecular Simulations of Ionic Liquid-Based Systems: From Structural to Optoelectronic Properties
Explore molecular simulations of ionic liquid systems, examining structural and optoelectronic properties for advanced applications.
A Large Deviation Perspective on Nanoscale Transport Phenomena
Explore nanoscale transport phenomena through large deviation theory, examining current fluctuations in nonequilibrium states and their relation to nonlinear responses in complex systems.
MaZe: A New Approach for Simulating Adiabatic Systems - Lecture
Explore MaZe, a novel approach for simulating adiabatic systems in physics. Learn about its applications in first principles simulations, polarizable systems, and ionic transport studies.
Force Spectroscopy Sheds Light on the Structure and Dynamics of Nucleosomes
Explore force spectroscopy's insights into nucleosome structure and dynamics, revealing chromatin's complex behavior at the molecular level.
From Thermal to Athermal Systems: Self-Assembly and Absorbing Phase Transitions
Explore self-assembly and absorbing phase transitions in thermal and athermal systems, bridging physics and biology in non-equilibrium processes.
Plastic Deformation of Superionic Water Ices
Explore plastic deformation in superionic water ices, examining unique properties and implications for planetary interiors and materials science.
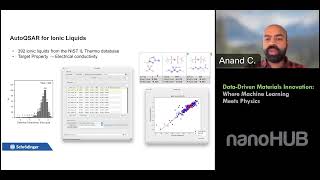
Data-Driven Materials Innovation: Where Machine Learning Meets Physics
Explore how machine learning and physics-based methods synergize to revolutionize materials science, addressing challenges in data-driven design across various technology fields with Schrödinger's innovative tools.