Benchmark and Critical Evaluation for ML Force Fields with Molecular Simulations
Explore benchmarking and critical evaluation of machine learning force fields in molecular simulations, focusing on realistic MD trajectories beyond force prediction accuracy.
ProtMD: Incorporating Conformation Flexibility for Drug Binding via Pretraining on MD Simulations
Explore ProtMD, a novel approach incorporating protein flexibility in drug binding predictions using molecular dynamics simulations and self-supervised learning techniques for improved accuracy.
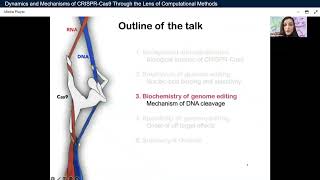
Dynamics and Mechanisms of CRISPR-Cas9 Through the Lens of Computational Methods
Explore computational methods revealing CRISPR-Cas9 dynamics and mechanisms, from RNA binding to large-scale conformational changes, enhancing understanding of this revolutionary genome-editing tool.
Scalable Molecular Simulation of Electrolyte Solutions with Quantum Chemical Accuracy
Explore advanced molecular simulation techniques for electrolyte solutions using neural network potentials and DFT, achieving quantum chemical accuracy and revealing novel insights in solution behavior.
Stability-Aware Boltzmann Estimator Training of Neural Network Interatomic Potentials
Explore advanced neural network interatomic potentials for molecular dynamics simulations using Stability-Aware Boltzmann Estimator Training to enhance stability and accuracy across various systems and architectures.
Молекулярна динаміка для фазового переходу рідина-газ - Частина 2
Delve into молекулярну динаміку та фізичні процеси фазового переходу рідина-газ через поглиблений теоретичний аналіз та сучасні дослідження.
Молекулярна динаміка для фазового переходу рідина-газ
Дослідіть молекулярну динаміку та механізми фазового переходу між рідким і газоподібним станами речовини через поглиблений теоретичний аналіз та моделювання.
Невідомий, Відомий Броунівський Рух
Дослідіть глибинні аспекти броунівського руху, від класичних концепцій до сучасних відкриттів, розкриваючи маловідомі властивості та практичне застосування цього фізичного явища.
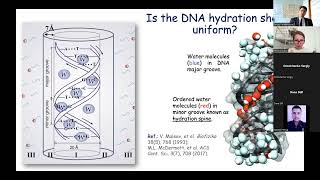
Молекулярно-динамічні моделювання колективної динаміки іон-гідратної оболонки подвійної спіралі ДНК
Дослідіть молекулярно-динамічне моделювання колективної поведінки іон-гідратної оболонки ДНК та її вплив на структурну динаміку подвійної спіралі.
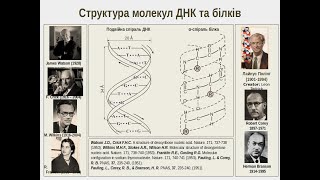
Метод класичної молекулярної динаміки в моделюванні властивостей молекули подвійної спіралі ДНК
Dive into classical molecular dynamics methods and their application in modeling DNA double helix properties, exploring computational approaches in modern biophysics research.
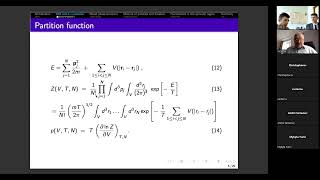
Молекулярна динаміка для фазового переходу рідина-газ - Частина 2
Дослідіть молекулярну динаміку та механізми фазового переходу між рідким і газоподібним станами речовини через поглиблений теоретичний аналіз та моделювання.
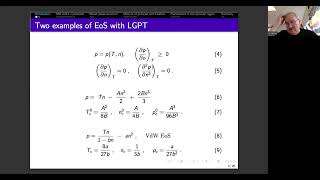
Молекулярна динаміка для фазового переходу рідина-газ
Дослідіть молекулярну динаміку та механізми фазового переходу між рідким і газоподібним станами речовини через поглиблений теоретичний аналіз та сучасні наукові підходи.
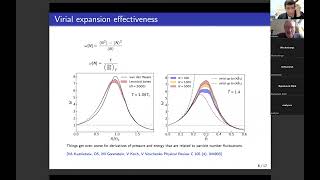
Флуктуації та Кореляції Частинок у Молекулярній Динаміці
Дослідіть флуктуації та кореляції частинок у системах з потенціалом Ленарда-Джонса, формування кластерів та їх вплив на метастабільні стани в молекулярній динаміці.
Estimating Normalizing Constants for Log-Concave Distributions - Algorithms and Lower Bounds
Explore algorithms and lower bounds for estimating normalizing constants in log-concave distributions, covering annealing, Monte Carlo methods, and Langevin dynamics.
Insights on Gradient-Based Algorithms in High-Dimensional Learning
Explore gradient-based algorithms in high-dimensional learning, covering statistical physics solutions for performance analysis in nonconvex settings like spiked mixed matrix-tensor model, perceptron, and phase retrieval.
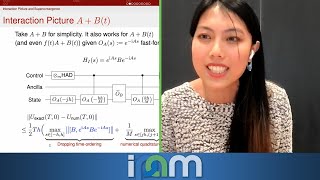
Time-Dependent Hamiltonian Simulation of Highly Oscillatory Dynamics - IPAM at UCLA
Explore highly oscillatory dynamics in quantum systems, focusing on efficient simulation algorithms for time-dependent Hamiltonians with applications in various scientific fields.
New Approaches in Simulation of Transition Paths - IPAM at UCLA
Explores tensor methods for simulating chemical transition processes, focusing on compressing high-dimensional PDE functions and introducing a novel tensor-network generative modeling approach without optimization.
Machine Learning and AI for the Sciences - Towards Understanding
Explore ML and AI applications in sciences, focusing on neuroscience, medicine, and physics. Learn about extracting insights from ML models to enhance understanding in fields like quantum chemistry.
Deep Learning and Numerical Methods Intersections for Improving Molecular and Fluid Dynamics
Explore intersections of deep learning and numerical methods to enhance molecular and fluid dynamics simulations, improving accuracy and efficiency in scientific computations.
Learning ML Interatomic Potentials - Day 2
Explore ML interatomic potentials for drug discovery with expert Gianni De Fabritiis. Gain insights into advanced techniques and applications in computational chemistry.
Flow Annealed Importance Sampling Bootstrap - Machine Learning for Complex Distributions
Explore Flow Annealed Importance Sampling Bootstrap for approximating complex target distributions. Learn about normalizing flows, annealed importance sampling, and applications to molecular systems.