Molecular Representations for Drug Discovery
Explore molecular representations for drug discovery, covering QSAR modeling, latent chemistry representations, bioactivity modeling, and de-novo design with phenotypic screens.
Drug Discovery Generative AI Using Tensor Network GPT or BERT
Explore generative AI applications in drug discovery using GPT and BERT models, focusing on ProtGPT2 for protein sequence generation and ProtBert for protein structure analysis.
Can LLMs Help Researchers Automate Individual Disease Drug Design?
Explore how Large Language Models enhance drug discovery through molecular simulation, reaction prediction, and database interaction for individual disease treatments.
Myriad LLM Approaches to Drug Discovery Generative Artificial Intelligence
Explore diverse LLM approaches in drug discovery GenAI, empowering medicinal chemists to enhance the process through informed prompts and molecule generation techniques.
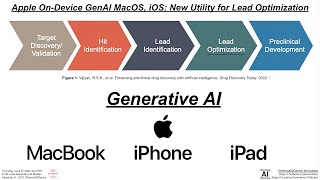
Apple On-Device GenAI for Lead Optimization in Drug Discovery - MacOS and iOS
Explore on-device GenAI capabilities for lead optimization in drug discovery using Apple devices. Learn about performance, speed, and memory for running AI models locally.
Chemical Foundation Models for Drug Discovery
Explore chemical foundation models for drug discovery, focusing on innovative approaches and potential breakthroughs in pharmaceutical research.
Is Medical Research Becoming Less Productive? - 28 August 2024
Explore the productivity trends in medical research with experts Nicholas Bloom and Maya Durvasula, examining challenges and potential solutions in the field.
Structured State-Space Sequence Models for De Novo Drug Design
Explore the potential of Structured State-Space Sequence (S4) models in advancing chemical language modeling for de novo drug design, including bioactive compound identification and natural product design.
Conditional Graph Information Bottleneck for Molecular Relational Learning
Explore a novel framework for molecular relational learning using graph neural networks to detect core subgraphs and predict interactions between molecular pairs.
Harmonic Self-Conditioned Flow Matching for Multi-Ligand Docking and Binding Site Design
Explore advanced AI techniques for protein-ligand binding structure generation and binding site design, improving drug discovery and molecular interaction modeling.
Large Language Molecular Representation and Learning
Explore molecular machine learning's potential in drug discovery, focusing on representation challenges, self-supervised learning, and leveraging pre-trained language models for property prediction.
PepFlow: Direct Conformational Sampling From Peptide Energy Landscapes
Explore PepFlow, a deep learning approach for direct conformational sampling of peptides. Learn about its modular design, hypernetwork integration, and applications in drug discovery.
Fostering the Development of Impactful AI Models in Drug Discovery - ICML 2023 Panel
Explore challenges and solutions in bridging academic AI research with real-world drug discovery applications through expert panel insights on benchmarking, data sharing, and industry-academia collaboration.
Binding Affinity Prediction with ML-Based Docking - Lab 2
Explore ML-based docking for binding affinity prediction in drug discovery, featuring insights from experts Stephan Thaler and Cristian Gabellini.
Protein Folding and Design - Day 5
Explore protein folding and design techniques with Alex Tong, delving into advanced machine learning applications for drug discovery and structural biology.
Target Deconvolution Explanation - Lab 4
Explore target deconvolution techniques in drug discovery through hands-on lab exercises led by experts Ali Denton and Kristina Ulicna.
Accelerate Atomistic Simulations, Sampling, and Dynamics - Day 2
Explore advanced techniques for accelerating atomistic simulations, enhancing sampling methods, and optimizing dynamics in drug discovery research.
Coarse Grained Biological Systems - Day 2
Explore coarse-grained biological systems in drug discovery with Jacopo Venturin, delving into advanced machine learning applications for pharmaceutical research.
Modeling Population Dynamics - Day 4
Explore population dynamics modeling techniques for drug discovery, focusing on innovative approaches and their applications in pharmaceutical research.
Open-Source Initiatives and Benchmarking Efforts in Machine Learning for Drug Discovery - Day 5
Explore open-source initiatives and benchmarking efforts in machine learning for drug discovery with Karmen Condic-Jurkic from Valence Labs.
Virtual Screening Techniques for Drug Discovery - Lab 1
Explore virtual screening techniques for drug discovery through hands-on lab exercises guided by industry experts.