AI-Driven Drug Design: Optimizing Pharmacokinetics and ADME
Explore AI-driven drug design techniques for optimizing pharmacokinetics and ADME with Novartis Principal Scientist Raquel Rodríguez Pérez at the Broad Institute symposium.
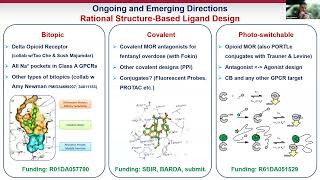
Computer-Driven Discovery of GPCR Ligands with New Chemotypes and Functional Selectivity
Explore computer-driven discovery of GPCR ligands, focusing on new chemotypes and functional selectivity. Learn structure-based techniques for rational ligand design and virtual screening.
ICM 3D Ligand Editor - Ligand Docking, Editing and Optimization
Explore ICM 3D Ligand Editor for advanced ligand docking, editing, and optimization techniques in molecular modeling and drug discovery.
Ligand Docking in ICM: Small Molecules, Fragments, Covalent and Template-Based Methods
Master ligand docking techniques in ICM-Pro, from pocket identification to covalent docking, enhancing your molecular modeling skills for drug discovery and design.
The Biology of Health Disparities: Ancestry-Driven Drug Discovery - K. Sean Kimbro
Explore ancestry-driven drug discovery and its potential to address health disparities, focusing on molecular characterization of hormonal cancers affecting individuals of African ancestry.
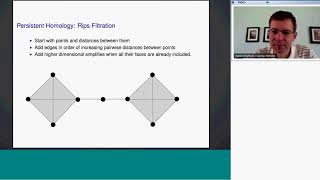
Anthony Bak - Spaces of Shapes Persistent Homology for Drug Discovery
Explore persistent homology in drug discovery, focusing on structure-based design, chemical compound analysis, and machine learning applications for virtual screening and classification.

Molecular Docking with AutoDock VINA - Script-Based Method for Multiple Ligands
Learn script-based molecular docking with AutoDock Vina for virtual screening of multiple ligands. Explore conformational space, calculate binding energies, and identify potential lead compounds efficiently.
Topological Data Analysis Based Machine Learning Models for Drug Design
Explores TDA-based models for effective molecular representation in drug design, combining persistent homology, spectral models, and Ricci curvature with machine learning for improved performance.
Methods for Computational Biology and Drug Discovery
Explore fundamental computational methods in physics and chemistry, presented by their creators, with insights into their development and applications.
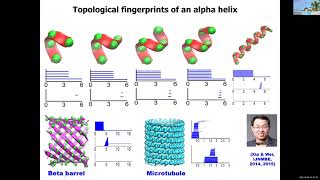
How Math and AI Are Revolutionizing Biosciences
Exploring how mathematics and AI revolutionize biosciences, focusing on tackling challenges in biological data complexity, dimensionality, and nonlinearity to enhance AI's capabilities in drug design and viral mutation prediction.
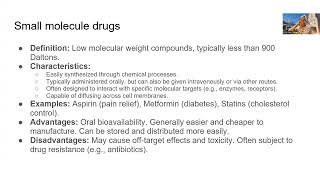
Introduction to Drug Development - Lecture 13
Explore the fundamentals of drug development, from initial research through clinical trials and regulatory approval processes in this comprehensive pharmaceutical sciences overview.
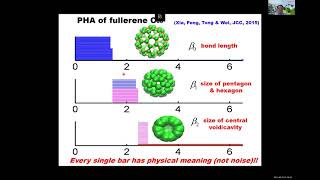
Kelin Xia - Persistent Function Based Machine Learning for Drug Design
Exploring persistent function-based machine learning for drug design, focusing on novel molecular representations and their application in improving predictive models for drug discovery.
Cyclica Recursion - Innovations in Drug Discovery
Explore cyclic recursion in machine learning for drug discovery, featuring innovative approaches and industry applications in pharmaceutical research.
Binding Affinity Prediction with ML-Based Docking - Lab 2
Explore ML-based docking for binding affinity prediction in drug discovery, featuring insights from experts Stephan Thaler and Cristian Gabellini.

Binding Affinity Prediction with Machine Learning-Based Docking - Lab 2
Explore ML-based docking techniques for predicting binding affinity in drug discovery, featuring insights from experts Stephan Thaler and Cristian Gabellini.
Lab 3 - De Novo Generation Explanation
Explore de novo molecule generation techniques with experts Emmanuel Bengio and Julien Roy in this recorded session from the 2024 Machine Learning for Drug Discovery Summer School.
Machine Learning in Structure-Based Drug Discovery - Day 2
Explore machine learning applications in structure-based drug discovery, focusing on innovative techniques and their impact on pharmaceutical research.
Leveraging Molecular ML and Property Prediction in Drug Design
Explore cutting-edge molecular machine learning techniques for drug design, focusing on property prediction to enhance pharmaceutical development.
Quantum Computing in the Cloud
Explore quantum computing's integration with classical HPC, its potential in solving complex problems, and Kvantify's innovative approach to combining quantum and classical computing technologies.
Quantum Computing in the Cloud
Explore quantum computing's integration with classical HPC, its potential in drug design, logistics, and finance, and Kvantify's innovative approach to solving complex computational problems.
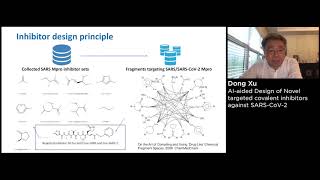
AI-Aided Design of Novel Inhibitors Against SARS-CoV-2
Explore AI-driven drug design for SARS-CoV-2, focusing on 3CLpro inhibitors. Learn about deep Q-learning networks, fragment-based design, and structure-based optimization for developing potential lead compounds.